
Metabolism
Abstract
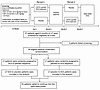
BACKGROUND Dichloroacetate (DCA) is an orally administered structural analog of pyruvate, an endogenous pyruvate dehydrogenase kinase inhibitor.METHODS We conducted a phase III multicenter trial in 34 children with pyruvate dehydrogenase complex deficiency (PDCD). Participants were randomly allocated to 4 months of treatment with DCA or a placebo, followed by a 1-month washout period and crossover to the alternate arm, and could continue into an open-label extension period. DCA dosing was predetermined by pharmacogenomic analysis of GSTZ1, which modulates DCA metabolism. The primary endpoint was the observer-reported outcomes motor domain (ObsROmotor) score. Additional assessments evaluated motor function, plasma lactate levels, and survival.RESULTS Chronic DCA was well tolerated and safe. The primary endpoint, ObsROmotor, was not statistically significantly different between the treatment and placebo groups (P = 0.512). However, longer-term treatment, including the open-label extension, showed a statistically significant treatment effect (P = 0.002), especially in participants with higher baseline motor impairment (ObsROmotor ≥ 8; P = 0.001). DCA decreased plasma lactate –0.48 (0.82) mmol/L (–20%; P = 0.006). Survival of participants was significantly greater than that of a natural history cohort (log-rank P = 0.027).CONCLUSION Longer-term treatment with DCA, dosed based on GSTZ1 haplotype, is safe and was associated with a statistically significant improvement in patient motor function, plasma lactate, and survival.FUNDING NIH (R01FD005407; R42HD089804), University of Florida Department of Medicine, Saol Therapeutics.
Authors
Peter W. Stacpoole, Jose E. Abdenur, Jirair K. Bedoyan, Lorenzo Botto, Gregory M. Enns, Marni J. Falk, Rebecca Ganetzky, Cheryl Garganta, Kevin Glinton, Andrea Gropman, Sharon Hamm, Eugenia Henry, Nicola Longo, Richard Neiberger, Russell P. Saneto, Fernando Scaglia, Sub H. Subramony, Jerry Vockley, Richard E. Wagner

Mitochonic acid 5 alleviates amyotrophic lateral sclerosis phenotypes via mitochondrial augmentation
Abstract
Amyotrophic lateral sclerosis (ALS) is a fatal neurodegenerative disease that urgently requires effective treatment. Mitochondrial dysfunction underlies ALS pathology and represents a potential therapeutic target. Here, we demonstrated the therapeutic potential of mitochonic acid 5 (MA-5), a novel mitochondria-targeted compound that ameliorated ALS phenotypes by enhancing mitochondrial function. In a Drosophila ALS model expressing a mutant human SOD1 (G85R), MA-5 significantly improved locomotor activity, with a trend toward restoration of mitochondrial integrity. In skin fibroblasts derived from ALS patients and motor neurons derived from induced pluripotent stem cells, MA-5 restored ATP production and increased mitochondrial motility. Multiomics analyses suggested that MA-5 modulated mitochondria-linked gene expression and downregulated the glycerophosphate shuttle, contributing to mitochondrial reactive oxygen species production. Transcriptomic analysis identified C7orf31 as a potential marker for monitoring the therapeutic effects of MA-5 and diagnosing ALS subtypes. These findings support MA-5 as a promising therapeutic candidate for ALS and propose C7orf31 as a potential biomarker for treatment monitoring and for disease subtyping.
Authors
Yoshitsugu Oikawa, Yuhan Luo, Naoki Suzuki, Tomoko Kasahara, Yoshiyasu Tongu, Yuki Yoshida, Tsukasa Tominari, Shogo Tanabe, Yoshiko Suto, Hitomi Kashiwagi, Saki Saito, Kensuke Ikeda, Chitose Suzuki, Arata Kuranaga, Tetsuya Akiyama, Satoru Morimoto, Yoshitsugu Aoki, Rieko Muramatsu, Tomoyoshi Soga, Masashi Aoki, Hideyuki Okano, Tetsuhiro Tanaka, Takaaki Abe, Erina Kuranaga, Takafumi Toyohara

Abstract
Foxp3 deficiency causes a profound loss of immune tolerance, unleashing autoreactive T and B cells, lymphoproliferation, cytokine-driven inflammation, and autoantibody production. This autoimmune pathology is fueled by increased glutamine usage, but it remains unresolved whether glutamine is necessary to produce energy or for intermediate metabolite biosynthesis responsible for immunomodulation. Here, we demonstrate that glutamine utilization for biosynthetic pathways supported autoimmune inflammation in the settings of Foxp3 deficiency and dextran sodium sulfate–induced colitis. By employing a model of autoimmunity driven by Treg-specific loss of Foxp3, we showed that this effect is independent of pathogenic Foxp3-deficient Treg reprogramming. Mechanistically, glutamine biosynthetic pathways sustained conventional T cell activation and proinflammatory cytokine production by preventing inosine accumulation and signaling, thus implicating adenosine pathway modulation in autoreactive T cell dysregulation. Conversely, autoreactive B cell activation and autoantibody production relied on glutamine-dependent asparagine availability, which we identified as a targetable vulnerability for autoantibody formation. These findings highlighted glutamine-driven biosynthetic processes as critical drivers of autoimmunity and revealed distinct metabolic vulnerabilities in autoreactive T and B cells that could be targeted for therapeutic intervention.
Authors
Mohammad Adeel Zafar, Charlotte N. Hill Machado, Jyotirmaya Behera, Yuelin Zhong, Xiao Li, Shakchhi Joshi, Yassine El Fazaa, Virginia Camacho, Peter Georgiev, Kiran Kurmi, Marcia Haigis, Louis-Marie Charbonnier
Abstract
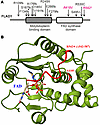
The FLAD1 gene codes for flavin adenine dinucleotide (FAD) synthase. FAD is a cofactor for many redox enzymes involved in vital processes from respiration to signal transduction. In this work, we described a clinical case of 2 siblings carrying compound heterozygous mutations in the FLAD1 gene resulting in the substitutions A418V and R542* at the protein level. The patients demonstrate adrenal insufficiency, which has not previously been associated with FLAD1 protein defects. To verify that adrenal insufficiency is caused by FLAD1 mutations, we created a personalized mouse model carrying the mutations found in the patients. The mutation in the FLAD1 gene, leading to the A418V substitution, appeared viable in the homozygous state, with minimal difference from the WT. The FLAD1 gene mutation leading to the R542* truncation is lethal when homozygous. The mouse model of the compound heterozygous FLAD1A418V/R542* mutations recapitulated the physiological, biochemical, and endocrine manifestations of FLAD1 mutations in patients. The mouse model created demonstrates the causal effect of FLAD1 mutations on the described pathology and potentially paves the way for understanding the disease’s molecular mechanism and developing better therapies.
Authors
Olga A. Averina, Natalia Yu. Kalinchenko, Vitaly A. Ioutsi, Andrey V. Pirogov, Alexander V. Bogachev, Oleg A. Permyakov, Vitaly S. Buev, Ekaterina A. Guseva, Anastasia V. Priymak, Olga A. Bazhanova, Mariia A. Emelianova, Olga O. Grigoryeva, Galina V. Baydakova, Maxim A. Abakumov, Vasily N. Manskikh, Olga A. Dontsova, Petr V. Sergiev, Anatoly N. Tiulpakov

Abstract
Wound healing is a highly dynamic and metabolically demanding process. However, the primary drivers of metabolic alterations involved in this process remain incompletely understood. Here, we employed multiomics profiling of clinical samples to investigate metabolic alterations during wound healing. Our analyses revealed significant activation of the TCA cycle and identified α-ketoglutarate (αKG) as a central regulator orchestrating the reparative phase. Systemic administration of αKG promoted wound closure and re-epithelialization, characterized by enhanced neo-tissue formation with an extended epithelial tongue. Mechanistically, αKG promoted cell proliferation via the cell cycle pathway and enhanced fibroblast-derived TGF-β signaling to induce epithelial-mesenchymal transition–like programs in epithelial cells. To address the spatial metabolic heterogeneity, we developed a transdermal MN platform based on gelatin methacryloyl for localized αKG delivery, further accelerating tissue repair. Collectively, these findings identify αKG as a metabolic driver of wound repair, reveal its dual role in modulating the epithelial-fibroblast microenvironment, and introduce a targeted bioengineering strategy with translational potential for both acute and chronic wound management.
Authors
Yuhan Li, Weimin Lin, Denghao Huang, Yueying Wang, Yimeng Cai, Jie Xiang, Linfeng Liu, Xinxing Shuai, Qi Yin, Shuang Jiang, Malcolm Xing, Yuan Wang, Leixiao Yu, Quan Yuan
Abstract
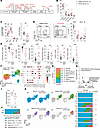
Gain-of-function (GOF) variants in STAT3 cause a complex disorder characterized by early-onset autoimmunity, lymphoproliferation, recurrent infections, and immune dysregulation. In both primary human and mouse models of STAT3 GOF, CD8+ T cells have been implicated as pathogenic drivers of autoimmunity, though the exact mechanisms remain poorly understood. Here, we found that in patients with STAT3 GOF, CD8+ T cells exist in an activated state. Functional assessment revealed that naive CD8+ T cells have an increased capacity for IFN-γ and TNF-α production, with type I and type II IFN transcriptional signatures. Evaluation of immunoregulatory pathways revealed dysregulation of the purinergic signaling axis in CD8+ T cells: CD39 was increased, whereas downstream purinergic family members, CD73 and the adenosine receptor A2AR, were downregulated, impairing the potential to produce or sense immunosuppressive adenosine. Evaluation of the impact of precision therapy, in the form of JAK inhibition, at a cellular and functional level revealed partial normalization of CD8+ T cell dysregulation in patients, including aberrant cytokine production. Our study suggests that a dysregulated purinergic signaling axis plays a key role in CD8+ T cell dysregulation in STAT3 GOF and may have implications for other rare monogenic immune disorders and common inflammatory disorders.
Authors
Jose S. Campos Duran, Montana S. Knight, Samir U. Sayed, Megan C. Dalalo, Andrea A. Mauracher, Peyton Conrey, Aaron B. Schultz, Ceire A. Hay, Robert B. Lindell, Ilona Neale, Kyle Yeakle, Eric D. Abrams, Erica G. Schmitt, Martin A. Thelin, Christian A. Howard, Sara Bluestein, Christine M. Seroogy, Tamara C. Pozos, Akaluck Thatayatikom, Ingrid S. Lundgren, Amelie Gauthier, Scott W. Canna, Helen C. Su, Michael D. Keller, Ottavia M. Delmonte, Lisa R. Forbes Satter, Steven M. Holland, Jenna R.E. Bergerson, Jennifer W. Leiding, Neil Romberg, Will Bailis, Christopher A. Hunter, Alexandra F. Freeman, Alejandro V. Villarino, Mark S. Anderson, Megan A. Cooper, Tiphanie P. Vogel, Sarah E. Henrickson
Abstract
Inflammatory cytokines reprogram keratinocyte metabolism, but the metabolic pathways that couple immune signals to pathological epidermal growth remain incompletely defined. Here, we identify GLS1-mediated glutaminolysis as a metabolic program preferentially induced in keratinocytes under type 3 inflammatory conditions. Integrated transcriptomic, metabolomic, genetic, and functional analyses showed that IL-17A induced GLS1 expression and glutaminolysis in keratinocytes. Keratinocyte-specific Gls1 deletion reduced the intracellular availability of arginine, proline, and methionine, impaired amino acid-dependent mTORC1 activation, disrupted redox homeostasis, and limited keratinocyte proliferation. Amino acid or antioxidant supplementation partially rescued these defects, whereas rapamycin blocked the amino acid-mediated proliferative rescue. Gls1 deletion did not impair steady-state skin development or homeostasis and did not alter MC903-induced type 2 dermatitis, but it delayed wound re-epithelialization and attenuated IMQ-induced psoriasiform inflammation. Loss of keratinocyte GLS1 also reduced epidermal chemokine expression and the accumulation of neutrophils and IL-17A-producing γδ T cells, revealing a role for glutaminolysis in amplifying epithelial-immune crosstalk. These findings define GLS1-mediated glutaminolysis as a context-specific metabolic checkpoint linking type 3 inflammation to keratinocyte proliferation and cutaneous immune amplification, and support locally or temporally controlled GLS1 inhibition as a potential therapeutic strategy for psoriasis.
Authors
Yongfei Hu, Hai Yu, Kaiyu Liang, Liyan Yuan, Wenjun Zhang, Zhe Zhuang, Suyun Ji, Xichun Xia
Abstract
Prediabetes associates with increased production of triglyceride-rich lipoproteins (TRLs), cardiovascular disease (CVD), and hepatic steatosis, which is linked to increased plasma levels of soluble TREM2 (sTREM2), the shed domain of TREM2 (triggering receptor expressed on myeloid cells 2). Whether and how TREM2 shedding contributes to elevated TRLs is unknown. By complementary analyses of individuals with prediabetes and hepatic steatosis and preclinical models, we show that plasma sTREM2 levels correlate positively with plasma apolipoprotein C3 (APOC3), an apolipoprotein that slows TRL catabolism and predicts CVD risk. Individuals with prediabetes and hepatic steatosis had higher plasma concentrations of APOC3-rich TRLs 35 to 60 nm in diameter than healthy controls. Mouse models of prediabetes with hepatic steatosis revealed that the increased plasma concentrations of sTREM2, APOC3, and TRLs were due to activation of macrophage ADAM17, a TREM2 sheddase. Preserving macrophage full-length TREM2 protected against the elevated plasma APOC3, sTREM2, dyslipidemia, and atherosclerosis, while TREM2-deficiency increased APOC3, TRLs, and atherosclerosis. Mechanistically, full-length TREM2 mediates macrophage TRL uptake, preventing excessive hepatic APOC3-rich TRL release and atherosclerosis. Our findings identify macrophage TREM2 shedding as an upstream contributor to the elevated TRLs in hepatic steatosis, providing a mechanistic link between hepatic steatosis and CVD risk in prediabetes.
Authors
Jingjing Tang, Jenny Kanter, Baohai Shao, Masami Shimizu-Albergine, Farah Kramer, Ah Reum Khang, Jason Luo, Huaqing Zheng, Alan Tran, Jocelyn Cervantes, Jeremy M Frey, Mauricio D. Dorfman, Cheng-Chieh Hsu, Laura J. den Hartigh, Tomas Vaisar, Brandon SJ Davies, Adam E. Mullick, George Ioannou, Gordon I Smith, Samuel Klein, Nicholas O. Davidson, Karin E. Bornfeldt

Abstract
Women with PMOS (formally termed PCOS) have an overall increased prevalence of metabolic syndrome (MetS) and central obesity. To help determine whether there might be changes in s.c. adipose tissue (SAT) associated with these abnormalities, we performed single-nuclei and scRNA-seq on SAT biopsies from 15 premenopausal PMOS women with signs of insulin resistance and 17 healthy BMI-matched controls. In SAT from PMOS versus control we observed a higher ratio of fibrotic versus insulin sensitive adipocytes and a higher ratio of mesenchymal stem cells (MSCs) to preadipocytes. Further in silico analysis suggested that preadipocytes in PMOS are more inflammatory and have a reduced capacity for differentiation. Slit homolog 2 (SLIT2), which is expressed at higher levels in MSC from PMOS, decreased adipogenesis in cell culture assays likely through its interaction with the Roundabout homolog 1 and homolog 2 (ROBO1/2) receptor expressed on the surface of preadipocytes. These new observations are consistent with higher SLIT/ROBO signaling, leading to reduced differentiation in the SAT of PMOS as an underlying mechanism for the aberrant ectopic fat accumulation and the development of MetS in PMOS.
Authors
Adeline Divoux, Edina Erdos, Katie L. Whytock, Timothy F. Osborne, Steven R. Smith

Abstract
Pulmonary alveolar proteinosis (PAP) is a rare pulmonary syndrome characterized by impaired surfactant clearance, driven by dysfunctional cholesterol efflux in alveolar macrophages (AMs). However, the molecular determinants governing AM cholesterol homeostasis remain incompletely defined. Here, through a genome-wide CRISPR screen in foamy macrophages and bulk RNA sequencing of AMs from PAP patients, we identify DTX4 as a pivotal regulator of cholesterol efflux in AMs. In mice, AAV-mediated silencing of DTX4 led to excessive AM lipid accumulation, exacerbated proteinosis, increased lung opacities, and deteriorated pulmonary function. Similarly, DTX4 depletion in primary AMs impaired cholesterol efflux and promoted intracellular lipid deposition. Conversely, AM-specific overexpression of DTX4 in the Csf2ra–/– PAP model markedly alleviated lipid accumulation, mitigated alveolar proteinosis, restored lung densities, and rescued pulmonary function. Mechanistically, DTX4 stabilizes the GM-CSF receptor via an E3-independent interaction to sustain JAK2/STAT5 signaling, which reciprocally maintains DTX4 transcription. This positive-feedback loop drives PPARγ expression, and its disruption in PAP impairs cholesterol efflux, a defect partially reversible by ectopic PPARγ expression. Collectively, our findings identify DTX4 as a central orchestrator of AM cholesterol efflux and surfactant homeostasis, positioning it as a promising therapeutic target for PAP.
Authors
Zimu Wang, Jingwei Shi, Xu Ye, Xinye Xia, Huihui Zhu, Qi Li, Min Chen, Yichao Zhao, Yingwei Zhang, Mengshu Cao, Yonglong Xiao, Xinmei Huang
No posts were found with this tag.


