

Abstract
Airway structural remodeling and hyperresponsiveness (AHR), hallmarks of asthma, are influenced by genetic variations and adverse exposures. While intrauterine perturbations in lung development have been linked to adult pulmonary disease, the developmental origins of these abnormalities remain poorly understood. Here, we provide evidence of genetic background playing a key role in this process. Using A/J and C57BL/6J mice known for their distinct susceptibility to AHR, we show that A/J embryos selectively develop an aberrant airway smooth muscle (SM) program and AHR in adulthood when exposed transiently to a vitamin A/retinoic acid (RA)–disrupted intrauterine environment in vivo by maternal BMS493 administration. Single-nucleus multiomics identified a mesenchymal cell population overactivating TGF-β targets in response to BMS493 selectively in A/J lungs. These cells, localized to sites of airway SM initiation and p-SMAD2- and -3, exhibited robust BMS493-mediated upregulation of SMAD2/3 targets, including regulators of SM program Pdgfra and Tnc. Functional analyses in vivo and cultured lungs showed aberrant SM formation in areas of overactive TGF-β of BMS493-exposed lungs. These abnormalities were prevented by inhibiting TGF-β signaling in utero in RA-deficient embryos. These findings underscore how distinct genetic backgrounds respond to intrauterine perturbations that program airway structure and function, with potential lasting consequences in postnatal pulmonary function.
Authors
Takehiro Otoshi, Benjamin D. Kotton, Ayyappa K.S. Kameshwar, Yoshinori Seki, Zachary Cardell, Xiangyi Ke, Yuta Matsuno, Pooja Rajaram, Youn-Kyung Kim, Sarah M. Sharpton, Loredana Quadro, Wellington V. Cardoso, Masako Suzuki

Abstract
Polycystic kidney disease (PKD) arises from mutations in cilia-associated genes, such as PKD1 and PKD2, expressed in renal epithelial cells, leading to progressive kidney dysfunction and end-stage kidney disease. Patients with PKD exhibit significant heterogeneity in disease progression, largely due to genetic and environmental modifiers. Like patients, mouse models of PKD also exhibit significant heterogeneity with regards to the gene mutated, age of disease onset, and rate of disease progression. To elucidate the cellular and molecular consequences of these variables, we constructed an integrated single-cell RNA sequencing (scRNA-seq) atlas across mouse models of PKD, mapping changes in cell type composition, gene expression, and intercellular signaling networks across the whole atlas and within individual models. Across models, scRNA-seq data revealed increased Spp1 (osteopontin) expression and signaling from PKD-enriched clusters. Global deletion of Spp1 in Pkd1RC/RC mice resulted in a modest reduction in cyst severity and improved kidney function. From these studies, we created a freely available, searchable website (https://bmblx.bmi.osumc.edu/scPKD/) that can be used to identify cross- and intra-model changes in gene expression, guiding researchers to new therapeutic targets for treating PKD.
Authors
Sarah J. Miller, Hua Zhong, Weidong Wu, Audrey M. Cordova, Morgan E. Yashchenko, Alex Yashchenko, Zhang Li, Daniyal J. Jafree, Chelsea N. Zimmerman, Christa I. DeVette, Vicki Do, Maya E. Hignite, Yohan Park, Fariha Nusrat, Bibi Maryam, Sizhao Lu, Xiaoyan Li, Jenny R. Gipson, Xiaogang Li, David A. Long, Mary C.M. Weiser-Evans, Bradley K. Yoder, Benjamin D. Cowley Jr, Katharina Hopp, Jason R. Stubbs, Qin Ma, Anjun Ma, Kurt A. Zimmerman

Abstract
Hypercapnia, elevated carbon dioxide (CO2), is common in advanced chronic obstructive pulmonary disease (COPD) and predicts poor clinical outcomes. Traditionally considered a consequence of disease severity, hypercapnia may drive disease progression by promoting airway dysfunction. Here, we show that hypercapnia acts as an active stressor, driving airway smooth muscle (ASM) constriction through a stromal interaction molecule 1–dependent (STIM1-dependent) pathway. Hypercapnia rapidly activates ERK, triggering sarcoplasmic reticulum calcium (Ca2+) release via phosphorylation of the inositol 1,4,5-trisphosphate receptor. ERK also induces nuclear translocation of the transcription factor c-Fos, enhancing STIM1 transcription. These responses were observed under both supraphysiological (~120 mmHg) and clinically relevant (50–60 mmHg) hypercapnia. Increased STIM1 abundance sustains store-operated Ca2+ entry (SOCE), amplifying ASM signaling. In mice, hypercapnia increased ASM and airway contractility in a STIM1-dependent manner. Human genetic analyses revealed noncoding STIM1 variants associated with reduced lung expression that were enriched in patients with COPD. These variants correlated with lower airway resistance under normocapnia; however, this benefit was lost during hypercapnia, indicating a potential gene-environment interaction. Together, our findings position STIM1 as a key mechanistic node linking hypercapnia to Ca2+ dysregulation and airway obstruction, defining a CO2/ERK/STIM1/SOCE axis with translational relevance to chronic lung disease.
Authors
Masahiko Shigemura, Vitalii Kryvenko, Jennifer A. Pacheco, Megan J. Puckelwartz, Milos Aleksic, Natalia D. Magnani, Emma Thompson, Francisco Javier Martin-Romero, Eoin P. Cummins, Werner Seeger, Andreas Bräuninger, Lynn C. Welch, G.R. Scott Budinger, Emilia Lecuona, Laura A. Dada, Ankit Bharat, István Vadász, Murali Prakriya, Jacob I. Sznajder

Abstract
Neonatal sepsis is a predominant cause of neonatal mortality and long-term morbidity that severely affects preterm and low-birth-weight newborns. Antibiotic resistance and long-term developmental issues associated with neonatal sepsis necessitate finding new and improved treatment options. IL-27 has diverse influences on the immune response, is elevated during the neonatal period compared with adulthood, and continues to rise further during infection. Elevated levels of IL-27 early in life predispose the host to impaired control of the pathogen burden and increased mortality. This study explored the therapeutic potential of IL-27p28 antibody administration to improve treatment outcomes during murine neonatal sepsis. Sepsis was induced by subcutaneous inoculation of K1-encapsulated E. coli, and the neonatal pups were rescued with IL-27p28 monoclonal antibody. Pups that received prophylactic antibody prior to the infection demonstrated superior bacterial clearance and significant weight gain compared with controls during infection. The combination of subclinical dose of gentamicin along with IL-27p28 antibody administered 2 hours after infection, significantly improved bacterial clearance and glucose homeostasis, with reduced serum levels of IL-6 and TNF-α, vital organ damage, and improved the survival rate of infected pups compared with gentamicin alone. These findings suggest that IL-27p28 antagonization represents a promising therapeutic tool for treatment of neonatal sepsis.
Authors
Madhavi Annamanedi, Jessica M. Povroznik, Samantha Arevalo-Marcano, Cory M. Robinson

Abstract
The pericardium plays an important homeostatic role for the neighboring heart, providing both lubrication and structural support. In vivo models have further identified a protective role for the pericardium in modulating cardiac remodeling following myocardial infarction, possibly through the actions of tissue-resident pericardial macrophages. Using patient-derived pericardial samples, we establish that human pericardial immune cells directly inhibit cardiac fibroblast fibrotic activity, and this action is dampened following myocardial infarction. Using single-cell RNA sequencing of patient pericardial fluid cells, we identify two pericardial macrophage subsets that are uniquely altered in response to myocardial infarction, which contributes to a shift in their effector molecule expression profiles. We confirm that fibronectin-expressing human pericardial macrophages are the primary driver of the pericardial antifibrotic actions through the release of cystatin C. Finally, we establish cystatin C as a myeloid cell–derived cardioprotective effector molecule in an in vivo model of myocardial infarction. Collectively, we uncover a molecular mechanism of the local immune environment that regulates cardiac remodeling after myocardial infarction.
Authors
Ali Fatehi Hassanabad, Sarthak Sinha, Arzina Jaffer, Darrell Belke, Nicole L. Rosin, Elodie Labit, Daniel Young, Friederike I. Schoettler, Keerthana Chockalingam, Benjamin Haeyul Lee, Jameson A. Dundas, Emilie de Chantal, Carmina A. Isidoro, Alexander Tam, Hanjoo B. Shim, Anna N. Zarzycki, Afshin Derakhshani, Elisabeth Gorgiogianni, Jeannine D. Turnbull, Antoine Dufour, Shalina S. Ousman, Jeff A. Biernaskie, Paul W.M. Fedak, Justin F. Deniset

Abstract
Locally produced nonpituitary growth hormone (npGH) promotes DNA damage accumulation and epithelial-mesenchymal transition (EMT) in aging human colon epithelium. GH receptor (GHR) and npGH are expressed in normal human prostate, and benign prostatic hyperplasia (BPH) prevalence increases with age. We hypothesized that local prostate GH action may promote EMT and contribute to BPH pathogenesis. We show here that the number of patients expressing npGH increases more than 10-fold after age 60, concordant with increased γH2AX, a marker of DNA damage, and EMT activation. GH-treated human primary prostate epithelial cells, normal prostate cells, and primary cell cultures derived from resected BPH specimens exhibited enhanced DNA damage and activated EMT, with induced TWIST2, suppressed E-cadherin, and increased Ki67, cell motility, and proliferation. In mice, prostate tissue adjacent to allografted GH-expressing fibroblasts showed increased γH2AX, TWIST2, and Ki67, along with morphological changes consistent with BPH. While GH and GH-induced IGF-1 both activated EMT, GH triggered DNA damage independently of IGF-1. These results elucidate what we believe to be a novel role for local npGH in aging prostate tissue, whereby npGH increases DNA damage and promotes EMT to enable a microenvironment favoring BPH development. Prostate GHR signaling may be an attractive therapeutic target for BPH.
Authors
Masaki Ryuzaki, Svetlana Zonis, Neil A. Bhowmick, Sandrine Billet, Saravana Kumar Kailasam Mani, Stephen J. Freedland, Hyung L. Kim, Vera Chesnokova, Shlomo Melmed

Abstract
Natural killer (NK) cells undergo stepwise differentiation from multipotent progenitors within secondary lymphoid tissues. Despite the central importance of the tissue microenvironment in their development, little is known about cell-cell interactions that regulate human NK cell trafficking and maturation. Here, we identify the chemokine receptor CXCR4 and its ligand CXCL12 as regulators of stromal–NK cell interactions required for NK cell maturation. We demonstrate that CXCR4 is expressed throughout human NK cell development in peripheral blood and tonsil, and CXCL12 is enriched in stromal niches containing developing NK cells. Pharmacologic blockade or genetic disruption of CXCR4 resulted in diminished adhesion to integrin ligands, and high-resolution imaging demonstrated crosstalk between CXCR4 and integrins, providing a mechanistic basis for chemokine-dependent modulation of adhesion. Further, CXCR4 blockade resulted in altered contact-dependent motility on stromal cells and integrin ligands, with decreased stable stromal engagement and increased cell speed. Consistent with a requirement for these interactions, treatment with the CXCR4 antagonist plerixafor (AMD3100) impaired NK cell generation from CD34+ precursors. Analysis of NK cells from patients with WHIM syndrome with CXCR4 gain-of-function mutations treated with plerixafor revealed similar defects in migration and adhesion, supporting the in vivo relevance of CXCR4-dependent regulation of NK cell adhesion and motility.
Authors
Shira E. Eisman, Francesca E. Grossberg, Batya S. Koenigsberg, David H. McDermott, Frédérique van den Haak, Luis A. Pedroza, Everardo Hegewisch-Solloa, Philip M. Murphy, Emily M. Mace
Abstract
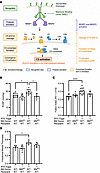
Schistosomiasis is a common cause of pulmonary hypertension (PH) worldwide. It is known that adaptive immunity and specifically CD4+ T cells are necessary for experimental disease pathogenesis. The lectin complement system is activated in those infected with schistosomiasis. We tested the hypothesis that lectin complement promotes Th2 CD4+ T cell activation, leading to PH in a schistosomiasis exposure model. WT and transgenic mice lacking mannose binding lectin (MBL), and bone marrow chimeras, were experimentally exposed to Schistosoma mansoni eggs. PH severity was assessed by hemodynamics and vascular remodeling, and CD4+ T cell density and phenotype were assessed by flow cytometry. WT recipients of MBL-knockout bone marrow were protected from Schistosoma-induced PH. The protection from PH was associated with fewer Th2 CD4+ T cells. In WT mice exposed to Schistosoma, CD4+ T cell expression of MBL increased. MBL-deficient CD4+ T cells had a suppressed Th2 phenotype when exposed to Schistosoma antigens. Mice with deficiency of C4, which functions downstream of MBL in the lectin complement pathway, were not protected from Schistosoma-induced PH. Mice lacking MBL were not protected from PH caused by hypoxia exposure. MBL in CD4+ T cells promotes Schistosoma-induced PH.
Authors
Claudia Mickael, Dara C. Fonseca Balladares, Rahul Kumar, Michael H. Lee, Kevin Nolan, Linda Sanders, Katie J. Tuscan, Ramraj Prasad, Pilar Londono, Fernanda P. Oliveira, Kennedi B. Pyper, Ari B. Molofsky, Rubin M. Tuder, Kurt R. Stenmark, Brian B. Graham
Abstract
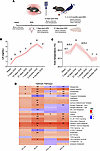
The hypothalamic changes that occur after the loss of ovarian estrogen remain poorly characterized. Here, we performed a comprehensive temporal characterization of the mouse hypothalamus after ovariectomy (OVX), combining physiological measurements with bulk RNA-seq of the posterior hypothalamus (PH) and preoptic area at 14 days and 4 months after OVX. Serum luteinizing hormone levels rose progressively and then declined, and core temperature peaked early and subsequently normalized, recapitulating the endocrine and thermoregulatory dynamics of reproductive aging in humans. Transcriptomic analysis revealed time-dependent activation of inflammatory pathways, glial markers, and KNDy neuron-related gene networks, with the most pronounced changes emerging at 4 months after OVX, particularly in the PH. Immunofluorescence confirmed increased neurokinin B release, declining KNDy neuronal activity, and heightened astrocytic reactivity in the arcuate nucleus after prolonged estrogen withdrawal. To contextualize these findings, we analyzed publicly available human hypothalamic RNA-seq data across chronological age. Age-related transcriptomic patterns, including progressive inflammatory signaling, glial activation, and altered KNDy gene expression, showed significant correlation with the OVX mouse model, particularly at the pathway level. These findings establish a temporal framework for hypothalamic molecular changes after estrogen withdrawal, identify conserved neuroinflammatory signatures across species, and provide a preclinical platform for testing interventions targeting menopause-associated hypothalamic dysfunction.
Authors
Jordana C.B. Bloom, Encarnación Torres, Sidney A. Pereira, Liliana Arvizu-Sanchez, Audrey N. Fontes, Hadine Joffe, David C. Page, Victor M. Navarro
Abstract
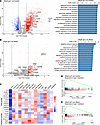
The precise mechanisms underlying the pathogenesis of idiopathic inflammatory myopathy (IIM) remain undefined. However, there has been increasing recognition that tissue-resident memory T cells (TRMs) play an important role in the pathogenesis of systemic autoimmune disease. In IIM, TRM-associated transcriptional signatures have been reported but on a very limited basis. By using multimodal single-cell RNA-sequencing analysis in our established murine model of histidyl-tRNA synthetase–induced myositis, we identified a prominent population of CD4+ TRMs in inflamed skeletal muscle. Muscle CD4+ TRMs exhibited high expression of genes encoding Cd69, Cxcr6, Runx3, and Prdm1, alongside low expression of Klf2, Ccr7, Sell, S1pr1, and Tcf7 — a profile that is generally consistent with previous reports of TRM gene signature and that we validate through comparison with transcriptomic profiles of human muscle tissue. Detailed pathway analysis in our model indicates that muscle CD4+ TRMs contribute to innate immune regulatory pathways enriched for TNF and IFN-γ signaling. Furthermore, analysis of TCR clonotype distribution and CDR3 sequence similarity revealed pronounced clonal expansion of CD4+ TRMs relative to other T cell subsets — a pattern that remained stable from 2 to 6 weeks after immunization. Collectively, these results suggest a potential role for CD4+ TRMs in the pathogenesis of autoimmune myositis.
Authors
Decheng Li, Daniel P. Reay, Iago Pinal-Fernandez, Maria Casal-Dominguez, Andrew L. Mammen, Sarah L. Gaffen, Timothy B. Oriss, Dana P. Ascherman
No posts were found with this tag.


